Sterilization Cycle Reproducibility is the ability of a validated sterilization process to consistently achieve its predefined critical process conditions and required sterilization effectiveness across repeated cycles. In pharmaceutical manufacturing, reproducibility is essential for maintaining sterility assurance and ensuring that products, equipment, components, and materials receive the required sterilization treatment.
A sterilization cycle may appear successful because the chamber reaches its programmed temperature and pressure. However, these readings alone may not demonstrate that every part of the load has received adequate sterilization treatment. Differences in load configuration, air removal, steam quality, equipment performance, initial load temperature, and sensor accuracy can influence cycle performance.
For example, an autoclave cycle used to sterilize laboratory media may show consistent chamber temperatures while certain densely packed items heat more slowly than others. If these differences are not understood during validation, some locations may receive less thermal exposure than intended.
Evaluating reproducibility involves reviewing repeated cycle data, heat distribution, heat penetration, critical process parameters, and relevant biological indicator results. A systematic approach helps pharmaceutical manufacturers identify variability, investigate failures, and maintain a validated state of control.
What Is Sterilization Cycle Reproducibility?
Sterilization cycle reproducibility refers to the consistency of sterilization process performance when the defined cycle is repeated under specified and controlled conditions.
A reproducible cycle should consistently meet its established acceptance criteria for the approved load configuration and operating range. These criteria are established during process development and validation.
Reproducibility does not mean that every recorded value must be numerically identical. Small variations can occur because of normal equipment and measurement variation. The important consideration is whether the observed variation remains within justified limits and whether the process continues to achieve its required sterilization effectiveness.
The evaluation should consider:
- Consistency of temperature and pressure profiles.
- Achievement of the specified exposure time.
- Uniformity of conditions throughout the sterilization chamber.
- Heat penetration into representative or worst-case load items.
- Required lethality, including F₀ where applicable to moist heat processes.
- Biological indicator results when used as part of the validation strategy.
- Equipment, monitoring system, and load configuration performance.
The applicable parameters depend on the sterilization method, the material being processed, and the approved validation strategy.
Why Is Sterilization Cycle Reproducibility Important?
1. Supports sterility assurance
Repeated successful cycles provide evidence that the sterilization process can consistently deliver the intended treatment. Reproducibility is one part of a broader sterility assurance strategy and does not, by itself, prove sterility of every individual unit.
2. Identifies process variability
Comparing cycle data helps reveal unexpected differences in heating, pressure, exposure time, or other critical parameters.
3. Supports batch quality decisions
When a cycle deviates from validated requirements, the recorded data and investigation findings help determine the potential impact on the load. A cycle should not be accepted solely because the final chamber temperature appears satisfactory.
4. Demonstrates control during validation
Validation establishes the evidence that the process can operate consistently under the defined conditions. Reproducibility studies help demonstrate that the selected cycle and load configuration are suitable.
5. Supports continued process verification
Routine cycle records and periodic review help identify deterioration in equipment performance, changes in load behaviour, and emerging trends that may require investigation or requalification.
Major Factors Affecting Sterilization Cycle Reproducibility
1. Load configuration
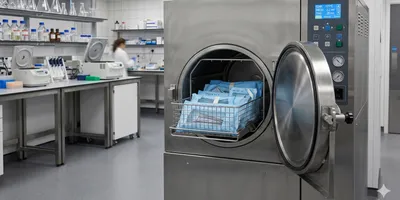
The arrangement, density, and quantity of items in a sterilizer influence heat transfer and the movement of steam or other sterilizing agents.
Overloading, tightly packed items, inappropriate wrapping, or changes in container arrangement may create locations that heat more slowly.
Control measure: Establish and document approved loading patterns. Evaluate relevant minimum and maximum loads and challenging configurations during validation.
2. Air removal and steam quality
For moist heat sterilization, inadequate air removal can interfere with effective steam contact and heat transfer. Non-condensable gases, unsuitable steam quality, and condensate accumulation may also affect process performance.
Control measure: Maintain the validated air removal process, monitor relevant steam quality attributes, and investigate recurring air removal or heating abnormalities.
3. Equipment performance
Faulty valves, vacuum system problems, steam supply fluctuations, worn seals, and control system issues can influence cycle performance.
Control measure: Maintain the sterilizer according to approved maintenance procedures, ensure calibration status is current, and investigate recurring alarms or abnormal trends.
4. Sensor accuracy and placement
Incorrectly calibrated sensors, poor sensor placement, and differences between control and independent monitoring systems can affect the interpretation of cycle results.
Control measure: Use suitable calibrated instruments and establish sensor locations through validation. Confirm that routine monitoring locations adequately represent the validated process.
5. Initial load conditions
The initial temperature, moisture content, and physical characteristics of a load can affect the time required to reach the required conditions.
Control measure: Define relevant preparation and loading requirements and evaluate challenging initial conditions where appropriate.
6. Operator practices
Differences in loading, wrapping, cycle selection, and adherence to approved procedures may introduce variability.
Control measure: Provide role-specific training and verify that operators consistently follow approved procedures.
How Is Sterilization Cycle Reproducibility Evaluated?
A structured evaluation combines process knowledge, physical measurements, and microbiological evidence where appropriate.
Step 1: Define the sterilization process
Identify the sterilizer, sterilization method, cycle recipe, material or product, load configuration, and intended use.
For moist heat sterilization, relevant parameters may include temperature, pressure, exposure time, air removal, and F₀ where applicable.
The process definition should specify critical parameters, acceptable ranges, monitoring requirements, and the approved load configurations.
Step 2: Develop a scientifically justified validation protocol
The protocol should define the study objective, test conditions, monitoring equipment, sensor locations, acceptance criteria, and data evaluation method.
The number of validation runs should be justified based on the process, applicable requirements, and quality risk assessment. Three consecutive successful cycles are commonly used in some qualification strategies, but three runs should not be treated as a universal requirement for every sterilization method or situation.
Step 3: Conduct heat distribution studies
Heat distribution studies evaluate how temperature is distributed throughout the sterilizer chamber.
Multiple calibrated temperature sensors are placed at justified locations to identify temperature differences and potential cold spots. Studies may also assess chamber performance under empty or defined loading conditions, depending on the purpose of the qualification.
The results help establish whether the chamber can deliver sufficiently uniform conditions under the evaluated circumstances.
Step 4: Conduct heat penetration studies
Heat penetration studies evaluate how quickly and effectively heat reaches the interior or most difficult-to-heat locations of the actual load.
For example, a densely filled container or a load containing wrapped materials may heat more slowly than an exposed metal component.
Temperature sensors are placed at justified locations within representative load items, including identified worst-case locations. The data help demonstrate that the required treatment is delivered throughout the load.
Step 5: Evaluate repeated cycle results
Compare the results from repeated cycles performed under the defined study conditions.
Review:
- Time required to reach the required temperature.
- Temperature distribution and minimum temperature achieved.
- Exposure time at the required conditions.
- Pressure and temperature relationships, where applicable.
- F₀ values or other validated lethality measures, where applicable.
- Differences between monitoring locations.
- Alarm events, aborted phases, or unusual fluctuations.
The assessment should demonstrate that the predefined acceptance criteria are met consistently. An average value should not be used to conceal an individual cycle or location that fails an applicable criterion.
Step 6: Assess biological indicators where applicable
Biological indicators may be used during sterilization validation to provide evidence of microbial inactivation under defined challenge conditions.
Their selection, placement, population, resistance characteristics, recovery method, and acceptance criteria should be justified for the sterilization process.
Biological indicator results complement physical measurements. They should not be used as a substitute for evaluating required physical process parameters.
Step 7: Document and approve the findings
Prepare a validation report summarising the protocol, load configurations, instrument details, raw data, deviations, results, acceptance criteria, and conclusion.
Any unexpected variation should be assessed before the process is declared acceptable. The report should be reviewed and approved through the site’s quality system.
Example: Evaluating Variability in an Autoclave Cycle
Consider a hypothetical autoclave validation study performed using the same approved load configuration across three consecutive runs.
| Parameter | Run 1 | Run 2 | Run 3 |
|---|---|---|---|
| Minimum load temperature during exposure | 121.2°C | 121.0°C | 121.1°C |
| Chamber pressure | Within the defined range | Within the defined range | Within the defined range |
| Exposure time | 20 minutes | 20 minutes | 20 minutes |
| Critical alarms | None | None | None |
| Acceptance criteria | Met | Met | Met |
These illustrative results suggest consistency in the recorded parameters. However, they do not independently establish that the cycle is suitable. The specified temperature, pressure, exposure duration, and required lethality must be justified for the particular process and load.
Now consider a separate run in which the chamber temperature reaches its set point, but a load sensor records a lower temperature at a challenging location.
This difference requires evaluation. Possible explanations include poor heat penetration, an unsuitable load arrangement, a sensor problem, or a steam distribution issue.
The run should not be declared acceptable solely because the chamber display reached its target. The applicable acceptance criteria and investigation findings must guide the decision.
Understanding Variability in Sterilization Cycles
Variability can be assessed by comparing individual measurements across repeated cycles and monitoring locations.
For numerical measurements, the following statistical concepts may help.
Mean
The mean represents the arithmetic average of a set of measurements.
[
\bar{x}=\frac{\sum_{i=1}^{n}x_i}{n}
]
Standard deviation
Standard deviation describes the spread of measurements around their mean.
[
s=\sqrt{\frac{\sum_{i=1}^{n}(x_i-\bar{x})^2}{n-1}}
]
A lower standard deviation generally indicates less observed variation within the dataset. However, low variation alone does not prove that a sterilization process is effective. All results must still meet the relevant acceptance criteria.
For small validation datasets, statistical conclusions should be interpreted cautiously. Three successful runs, for example, may provide evidence of repeatability under the tested conditions, but they cannot represent every possible source of variability.
Common Causes of Sterilization Cycle Failure
| Observation | Possible cause | Recommended investigation |
|---|---|---|
| Slow temperature rise | Steam supply or air removal problem | Review cycle trends, utility conditions, and air removal performance |
| Temperature differences between locations | Load arrangement or heat distribution issue | Review sensor placement and repeat relevant mapping studies if justified |
| Insufficient heating within a load item | Poor heat penetration or challenging load characteristics | Assess the load design and identify the slowest-heating locations |
| Unexpected pressure fluctuations | Valve, vacuum, or control system issue | Review alarms, pressure trends, maintenance, and calibration records |
| Inconsistent lethality calculations | Temperature variability, sensor errors, or calculation issues | Verify source data, calculation settings, sensor calibration, and process assumptions |
| Positive biological indicator | Inadequate treatment, contamination, or a test-related issue | Investigate the entire process, biological indicator handling, and recovery controls |
These are possible causes, not automatic conclusions. The investigation should establish the actual or most probable cause using documented evidence.
How Should Variability Be Investigated?
When a sterilization cycle produces unexpected results, the investigation should begin by establishing what happened and which acceptance criteria were affected.
A practical approach includes:
- Review cycle records: Examine temperature, pressure, exposure time, alarms, and relevant process trends.
- Verify the load: Confirm that the approved load pattern, wrapping, quantity, and cycle recipe were used.
- Check monitoring equipment: Review calibration status, sensor placement, and instrument performance.
- Evaluate utilities and equipment: Investigate steam quality, air removal, valves, vacuum performance, and maintenance history as relevant.
- Assess the load impact: Determine whether the deviation could affect the required sterilization treatment and identify all potentially affected items or batches.
- Establish the cause: Use appropriate root cause analysis rather than assuming operator error or equipment failure without evidence.
- Define corrective actions: Implement actions proportionate to the confirmed cause and assess their effectiveness.
- Determine the need for requalification: Significant changes or unresolved performance concerns may require additional qualification or validation work before routine use resumes.
Affected loads should remain under appropriate control until an authorised quality decision is made. Repeating the cycle or adding extra exposure must not be used as an automatic correction without an approved, scientifically justified procedure.
Requalification and Ongoing Process Verification
Sterilization validation is not a one-time activity. The validated state must be maintained through routine monitoring, equipment maintenance, calibration, deviation management, and periodic review.
Requalification or revalidation may be required following changes or events that could affect sterilization effectiveness, such as:
- Major equipment repair or modification.
- Changes to cycle parameters or control software.
- Changes to load configuration or packaging.
- Changes in the material or product being sterilized.
- Repeated cycle failures or unexplained variability.
- Evidence that the validated worst-case conditions are no longer adequately represented.
The scope and timing should follow applicable requirements, the approved validation programme, and a documented risk assessment.
WHO GMP guidance for sterile pharmaceutical products recommends periodic review and verification of sterilization processes. For heat sterilization, it specifies at least annual revalidation for load patterns considered worst case, with the frequency for other load patterns justified through the contamination control strategy. Applicable regional requirements should be checked for the facility concerned.
Common Mistakes to Avoid
- Relying only on chamber temperature: This may not demonstrate adequate heat penetration into the load.
- Ignoring the worst-case load: A cycle that works for a small, simple load may not work equally well for a dense or complex load.
- Using an unjustified number of runs: Validation study design should be scientifically justified and risk-based.
- Treating biological indicators as the only evidence: Physical measurements and other required process controls remain important.
- Using average results to hide failures: Individual acceptance criteria and challenging locations must be assessed.
- Changing the loading pattern without evaluation: Load changes can alter heat transfer and process performance.
- Failing to trend routine cycle data: Gradual deterioration may remain unnoticed without periodic review.
Frequently Asked Questions
1. What is sterilization cycle reproducibility?
It is the ability of a sterilization process to consistently meet its predefined process requirements and demonstrate the intended sterilization effectiveness across repeated cycles.
2. How many cycles are needed to demonstrate reproducibility?
The number depends on the sterilization method, validation strategy, applicable requirements, and risk assessment. Three consecutive successful runs are used in some qualification approaches, but this is not a universal rule for all processes.
3. What is the difference between heat distribution and heat penetration?
Heat distribution evaluates temperature uniformity throughout the sterilizer chamber. Heat penetration evaluates how heat reaches the interior or most difficult-to-heat locations of the actual load.
4. Can a cycle pass if the chamber temperature is acceptable but one load sensor fails?
Not automatically. The failed or unexpected reading must be investigated against the validated acceptance criteria. A satisfactory chamber display alone does not establish that the entire load received adequate treatment.
5. Do biological indicators prove that every item is sterile?
No. Biological indicators provide evidence of microbial inactivation at the tested challenge locations under the conditions of the study. Their results must be interpreted alongside physical process data and the overall validation strategy.
6. When should sterilization cycle reproducibility be reassessed?
Reassessment may be needed after significant equipment, process, product, packaging, or load changes; recurring failures; or evidence of unexplained variability. Periodic review should also follow the applicable validation programme and regulatory expectations.
Conclusion
Sterilization Cycle Reproducibility is a fundamental element of pharmaceutical sterilization validation. It demonstrates that a defined process can consistently achieve its required conditions across repeated cycles and approved load configurations.
Reliable evaluation requires more than reviewing chamber readings. Heat distribution, heat penetration, critical process parameters, equipment performance, load configuration, and biological indicator evidence where applicable must be assessed together.
By using a scientifically justified validation protocol, investigating unexpected variation, maintaining complete records, and performing appropriate periodic reviews, pharmaceutical manufacturers can strengthen process control and support reliable sterility assurance.
References
- World Health Organization (WHO). WHO Technical Report Series No. 1044, Annex 2: WHO Good Manufacturing Practices for Sterile Pharmaceutical Products.
https://www.who.int/docs/default-source/medicines/norms-and-standards/guidelines/production/trs1044-annex-2-gmp-for-sterile-pharmaceutical-products.pdf - Health Canada. Guide to Validation of Terminal Sterilization Process of Drugs (GUI-0074): Overview.
https://www.canada.ca/en/health-canada/services/drugs-health-products/compliance-enforcement/good-manufacturing-practices/guidance-documents/guide-validation-terminal-sterilization-process-drugs-gui-0074.html - Health Canada. Process Validation, Sterilization by Moist Heat: Guide to Validation of Terminal Sterilization Process of Drugs.
https://www.canada.ca/en/health-canada/services/drugs-health-products/compliance-enforcement/good-manufacturing-practices/guidance-documents/guide-validation-terminal-sterilization-process-drugs-gui-0074/process-validation-sterilization-moist-heat.html - U.S. Food and Drug Administration (FDA). Sterilization Process Controls.
https://www.fda.gov/inspections-compliance-enforcement-and-criminal-investigations/inspection-guides/sterilization-process-controls
This article is for professional educational purposes. Always follow the applicable regulations, approved validation protocol, established acceptance criteria, and site-specific procedures.